Cleancap mRNA Invitro Transcription: Difference between revisions
| (2 intermediate revisions by the same user not shown) | |||
| Line 48: | Line 48: | ||
[[File:ARCA Cap Analog.png|RTENOTITLE]] | [[File:ARCA Cap Analog.png|RTENOTITLE]] | ||
https://www.targetmol.com/compound/sb%20216641%20hydrochloride | |||
| Line 88: | Line 88: | ||
Pseudouridine (5-ribosyluracil) was the first modified ribonucleoside discovered. It is the most abundant natural modified RNA base, and has been deemed the "fifth nucleoside" in RNA. It can be found in structural RNAs, such as transfer, ribosomal and small nuclear RNA. Pseudouridine has been found to enhance base stacking and translation. Pseudouridine-5'-triphosphate (Pseudo-UTP) is used to impart desirable mRNA characteristics such as increased nuclease stability, increased translation or altered interaction of innate immune receptors with in vitro transcribed RNA. Pseudo-UTP, along with 5-methylcytidine-5'-triphosphate (5-methyl-CTP) has shown innate immune suppression in culture and in vivo while enhancing translation in recent publications. For example, Warren et al. determined an efficient means of reprogramming multiple human cell types using modified mRNA that can express the four primary reprogramming proteins. These cells are referred to as induced pluripotency stem cells (iPSCs). Warren et al. found that enzymatically synthesized RNA substituted with Pseudo-UTP, 5-Methyl-CTP and ARCA effectively evaded the cell’s innate immune response, a crucial component in their success. Reduced toxicity due to substitution with Pseudo-UTP, 5-Methyl-CTP was critical since it allowed repeated transfection with in vitro transcribed mRNA over several weeks. | Pseudouridine (5-ribosyluracil) was the first modified ribonucleoside discovered. It is the most abundant natural modified RNA base, and has been deemed the "fifth nucleoside" in RNA. It can be found in structural RNAs, such as transfer, ribosomal and small nuclear RNA. Pseudouridine has been found to enhance base stacking and translation. Pseudouridine-5'-triphosphate (Pseudo-UTP) is used to impart desirable mRNA characteristics such as increased nuclease stability, increased translation or altered interaction of innate immune receptors with in vitro transcribed RNA. Pseudo-UTP, along with 5-methylcytidine-5'-triphosphate (5-methyl-CTP) has shown innate immune suppression in culture and in vivo while enhancing translation in recent publications. For example, Warren et al. determined an efficient means of reprogramming multiple human cell types using modified mRNA that can express the four primary reprogramming proteins. These cells are referred to as induced pluripotency stem cells (iPSCs). Warren et al. found that enzymatically synthesized RNA substituted with Pseudo-UTP, 5-Methyl-CTP and ARCA effectively evaded the cell’s innate immune response, a crucial component in their success. Reduced toxicity due to substitution with Pseudo-UTP, 5-Methyl-CTP was critical since it allowed repeated transfection with in vitro transcribed mRNA over several weeks. | ||
https://www.targetmol.com/compound/mrtx-1719 | |||
[[File:Pseudouridine-5'-Triphosphate.png|400px|RTENOTITLE]] | [[File:Pseudouridine-5'-Triphosphate.png|400px|RTENOTITLE]] | ||
| Line 846: | Line 847: | ||
<span style="color: rgb(37, 37, 37); font-family: sans-serif; font-size: 14px;">Denaturing chromatographic purification<br/>The glass columns (890/10 mm) and (xk16/1000 mm) were packed according to manufacturer’s specifications with gel filtration matrix (Superdex-75, Superdex-200) and equilibrated in a cold room (4°C) with several column volumes (CV) of specified buffer. The transcription mixture from the previous step was loaded directly onto the equilibrated SEC column, and the chromatography was performed at 1 mL/min flow rate, collecting 2 mL fractions. Fractions were analyzed by denaturing urea-PAGE. S-box RNA containing fractions were combined and concentrated to 0.5 mL using Millipore centrifugal filter units (EMD Millipore), washed twice with the appropriate buffer for chemical probing analysis. All experiments were performed at 4°C. The efficiency of proposed purification system was confirmed through SHAPE experiments for S-box RNA.</span> | <span style="color: rgb(37, 37, 37); font-family: sans-serif; font-size: 14px;">Denaturing chromatographic purification<br/>The glass columns (890/10 mm) and (xk16/1000 mm) were packed according to manufacturer’s specifications with gel filtration matrix (Superdex-75, Superdex-200) and equilibrated in a cold room (4°C) with several column volumes (CV) of specified buffer. The transcription mixture from the previous step was loaded directly onto the equilibrated SEC column, and the chromatography was performed at 1 mL/min flow rate, collecting 2 mL fractions. Fractions were analyzed by denaturing urea-PAGE. S-box RNA containing fractions were combined and concentrated to 0.5 mL using Millipore centrifugal filter units (EMD Millipore), washed twice with the appropriate buffer for chemical probing analysis. All experiments were performed at 4°C. The efficiency of proposed purification system was confirmed through SHAPE experiments for S-box RNA.</span> | ||
https://www.targetmol.com/compound/mrtx-1719 | |||
== <span style="color: rgb(37, 37, 37); font-family: sans-serif; font-size: 14px;">Transfection</span> == | == <span style="color: rgb(37, 37, 37); font-family: sans-serif; font-size: 14px;">Transfection</span> == | ||
| Line 995: | Line 998: | ||
Mlu1<br/>acgcgT CGAGGGGAATTAATTC<br/> (VEE-pA-R GAATTAATTCCCCTCGAcgcgttctag ) | Mlu1<br/>acgcgT CGAGGGGAATTAATTC<br/> (VEE-pA-R GAATTAATTCCCCTCGAcgcgttctag ) | ||
[[Category:Pages with broken file links]] | [[Category:Pages with broken file links]] | ||
https://www.targetmol.com/compound/mrtx-1719 | |||
== EGFP mRNA Protocol == | == EGFP mRNA Protocol == | ||
Latest revision as of 01:09, 6 March 2026
Protocol for Co-transcriptional capping using CleanCap® Reagent AG from TriLink and HiScribe™ T7 High Yield RNA Synthesis Kit from New England Biolabs®
[edit]Reagents required:
1. NEB HiScribe™ T7 High Yield RNA Synthesis Kit, NEB # https://www.neb.com/products/e2040-hiscribe-t7-high-yield-rna-synthesis-kit E2040S
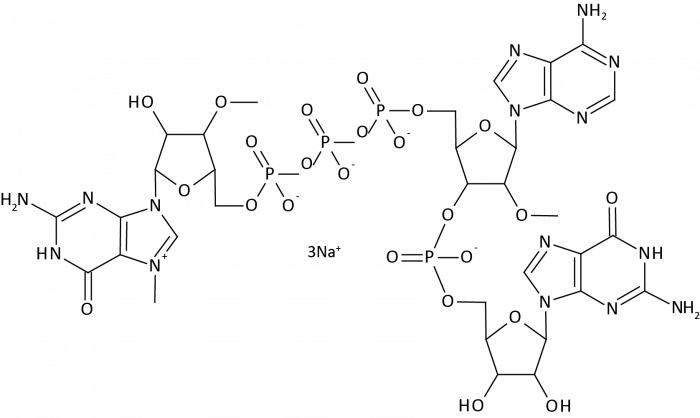
2. TriLink CleanCap® Reagent AG, N-7113, or CleanCap® Reagent AG (3’ OMe), N-7413
| Extinction Coefficient | 30,539 Lmol-1cm-1 at 255 nm |
|---|---|
| Molecular Formula | C33H45N15O24P4 (free acid) |
| Molecular Weight | 1159.60 |
| Salt Form | Na+ |
| Concentration | 100 mM |
| Buffer | H2O |
| Recommended Storage | -20°C or below |
| Other Name(s) | CleanCap®Reagent AG (3' OMe) for co-transcriptional capping of mRNA, m7(3'OMeG)(5')ppp(5')(2'OMeA)pG |
10 nmole ==>11.6 mg
to make 100 mM, 11.6 mg + 100 ul H2O
to calculate more acculately, dilute 1000 fold, use nanodrop, 30,539 Lmol-1cm-1 at 255 nm
ARCA (m27,3'-OGP3G or 3′-O-Me-m7G(5′) ppp(5′)G
[edit]
https://www.targetmol.com/compound/sb%20216641%20hydrochloride
CleanCap AG (3'OMe)
[edit]
Cap analogues
[edit]

Pfizer

Trinucleotide cap1 analogue
[edit]((m2 7,3’-O)Gppp(m2’-O)ApG; TriLink)
UTP
[edit]Uridine 5'-Triphosphate, Sigma , 94370-1G, 1 g $288

Pseudouridine-5'-Triphosphate, Pseudo-UTP, 5-Ribosyl Uracil, N-1019-5,5 µmole $451
Pseudouridine (5-ribosyluracil) was the first modified ribonucleoside discovered. It is the most abundant natural modified RNA base, and has been deemed the "fifth nucleoside" in RNA. It can be found in structural RNAs, such as transfer, ribosomal and small nuclear RNA. Pseudouridine has been found to enhance base stacking and translation. Pseudouridine-5'-triphosphate (Pseudo-UTP) is used to impart desirable mRNA characteristics such as increased nuclease stability, increased translation or altered interaction of innate immune receptors with in vitro transcribed RNA. Pseudo-UTP, along with 5-methylcytidine-5'-triphosphate (5-methyl-CTP) has shown innate immune suppression in culture and in vivo while enhancing translation in recent publications. For example, Warren et al. determined an efficient means of reprogramming multiple human cell types using modified mRNA that can express the four primary reprogramming proteins. These cells are referred to as induced pluripotency stem cells (iPSCs). Warren et al. found that enzymatically synthesized RNA substituted with Pseudo-UTP, 5-Methyl-CTP and ARCA effectively evaded the cell’s innate immune response, a crucial component in their success. Reduced toxicity due to substitution with Pseudo-UTP, 5-Methyl-CTP was critical since it allowed repeated transfection with in vitro transcribed mRNA over several weeks.
https://www.targetmol.com/compound/mrtx-1719

5-Methyluridine-5'-Triphosphate , 5-Methyl-UTP, 5-Me-UTP, 5-mUTP, Trilink: N-1024-5,5 µmole $464.
Thermostability Enhancer

N1-Methyl-Pseudouridine-5'-Triphosphate, N1-Methylpseudo-UTP, 1-Methylpseudo-UTP, Trilink: N-1081-5,5 µmole $464
Incorporation of N1-methylpseudouridine can reduce the immunogenicity of the resulting mRNA

https://www.targetmol.com/compound/uridine_5_triphosphate_tetrasodium
Reagents not supplied:
RiboMAX™ Large Scale RNA Production Systems – T7: Promega P1300
ScriptCap™ m7G Capping System: CellScript C-SCCE0625
ScriptCap™ 2'-O-Methyltransferase Kit: CellScript C-SCMT0625
A-Plus™ Poly(A) Polymerase Tailing Kit: CellScript C-PAP5104H
Antarctic Phosphatase: New England BioLabs M0289S
ScriptGuard™ RNase Inhibitor: CellScript C-SRI6310K
Lipofectamine™ MessengerMAX™ Transfection Reagent: ThermoFisher LMRNA001
RiboJuice™ mRNA Transfection Kit: MilliporeSigma TR-1013
Design of Transgene into T7-VEE
[edit]1, add Kozak after Nde1
Nde1 Kozak start
catatg gaa t tcc gcc acc ATG---
2, Add stop codon and Kozak after Nde1
Nde1 stop Kozak start
catatg agg ttc tag gcc acc ATG
                          -17                   +1-->
T7 promoter class III 5'- TAA TAC GAC TCA CTA TA G GGA GAC ATG CTA - 3'
T7 promoter class II  5'- TAA TAC GAC TCA CTA TT A GGG AGA ATG CTA - 3'

T7 RNA Polymerase
+ rNTPs
--->  RNA 5'- pppGGGAGACAUGCUA
5'UTR
[edit]1, T7 + beta globin + kozak
TAATACGACTCACTATAaGG GAATACAAGCTACTTGTTCTTTTTGCA AATTGCCACCATG
2, T7 + Venezuelan equine encephalitis virus
TAATACGACTCACTATAaGG ATGGGCGGCGCATGAGAGAAGCCCAGACCAATTACCTACCCAAA
Template preparation
[edit]1. Double-stranded DNA template
**NOTE: CleanCap AG requires modification to the sequence just downstream of the T7 promoter sequence, replacing the “GG” that follows the T7 promoter sequence with an “AG” as follows:
Standard T7 promoter (underlined) with 'GG':
5’ TAATACGACTCACTATA'GG' 3’
Sequence required for CleanCap® with AG:
5’ TAATACGACTCACTATA'AG' 3’
New primer
T7-mRNA-F (for pCDNA5 plasmid) (Box O1well I3)
AAATTAATACGACTCACTATAAG
T7-mRNA-F2 (for pCDNA5 pCMV5-frt plasmid)
ATTAATACGACTCACTATAAGGAGACCCAAG
T7-mRNA-F3 (for plasmid with traditional T7 promoter and kozak seq, such as pMiniT2--IRES)
TAATACGACTCACTATAAGGAGG
T7-mRNA-F4 (for plasmid vector pCAGGS, 2724)
TAATACGACTCACTATAAGGCAAGGCCACCGCCACCATG
Reverse primer for 2724 pCAGGS: pCAGGS-mRNA-R2
tTTTTTTTTTTTTTTTTTTTTTTTAAATGATTTGCCCTCC
(GGAGGGCAAATCATTTAAAAAAAAAAAAAAAAAAAAAAAa)
reverse primer for C-Flag tag
Flag-R
(CAAAGACGATGACGACAAGTAACTCGAGAAAAAAAAAAAAAAA)
TTTTTTTTTTTTTTTCTCGAGTTACTTGTCGTCATCGTCTTTG
2. NEB Monarch® RNA Cleanup Kit (500 µg), https://www.neb.com/products/t2050-monarch-rna-cleanup-kit-500-ug T2050
For pMiniT2-pA vector pA-T7-F
TAATACGACTCACTATAGGGAGAC
pA-mRNA-R
TAGGTAATCGTCACCTGcag
(ctgCAGGTGACGATTACCTA)
Co-transcriptional Capping Protocol:
(NOTE: 5 mM final each NTP instead of 10 mM in standard HiScribe Reaction)
- Thaw the necessary components, keep the T7 RNA Polymerase Mix on ice.
- Mix and pulse-spin in a microfuge to collect the solutions to the bottom of the tubes.
- Set up the reaction at room temperature in the following order:
|
Component |
Volume |
Concentration |
|---|---|---|
|
Nuclease-free Water |
X µl |
|
|
10X Reaction Buffer |
2 µl |
0.5X final |
|
100 mM ATP |
2 µl |
5 mM final |
|
100 mM GTP |
2 µl |
5 mM final |
|
100 mM pseudo-UTP |
2 µl |
5 mM final |
|
100 mM 5m-CTP |
2 µl |
5 mM final |
| Cyanine 5-UTP, optional |
0.5 µl | |
|
100 mM CleanCap'®' AG or AG (3’ OMe) (N-7113 or N-7413) |
1.6 µl |
4 mM final |
|
Linear Template DNA |
X µl |
1 µg total |
|
T7 RNA Polymerase Mix |
2 µl |
|
|
Total |
40 µl |
|
1.Mix thoroughly by pipet and pulse-spin in a microfuge. Incubate at 37°C for 2 hours in a dry air incubator or PCR machine to prevent evaporation.
2. Follow the protocol provided with the HiScribe T7 High Yield RNA Synthesis Kit (NEB E2040S ) for DNase treatment and RNA purification.
3. Add TURBO DNase (ThermoFisher Scientific, incubated at 37 ◦C for 30 min Purify with column
4, Add 5 µL of 10×Phosphatase Buffer to the reaction.
5, Add 1 µL of RNase-free DNase
6, Add 2 µL Antarctic Phosphatase (New England Biolabs) inccubate 37 ◦C for 30 min ( Phosphatase treatment reduces innate immune response in mammalian cells)
'5m-CTP' : 5-Methylcytidine-5'-Triphosphate
'pseudo-UTP': Pseudouridine-5'-Triphosphate
Cyanine 5-UTP, 100 nmol NEL583001EA $646.00 , ENZ-42506, 250 nmol, $354.00 , sc-499469, 250 nmol $340
Cyanine 3-UTP, 100 nmol NEL582001EA $641 25ul (10mM) $195 sc-499471 250 nmol $350
Large scale production
[edit]CleanCap AG (TriLink N-7113) and RiboMAX™ Large Scale RNA Production Systems-T7 (Promega P1300)
| Component |
Large Scale | Mid Scale | Small scale |
| T7 Transcription Buffer 5X |
20 μL |
||
| 100 mM ATP |
7.5 μL | ||
| 100 mM GTP |
7.5 μL |
||
| 100 mM 5mCTP |
7.5 μL |
||
| 100 mM pseudo-UTP |
7.5 μL |
||
| CleanCap AG, 100 mM |
7.5 μL |
||
| DNA plasmid (1 μg/μL) |
10 μL |
||
| RNAse-free water | 22.5 μL | ||
| Enzyme Mix (T7) |
10 μL |
||
| Total | 100 μL |
||
| Incubate at 37°C for 2 hrs. Do not incubate beyond 2 hours at 37°C as a longer incubation period may result in increased RNA degradation. | |||
| Add 10 μL RQ1 RNase-free DNase (Promega M6101) for 10 μg of template DNA (use 1 μL per μg DNA template). Gently mix by pipetting up and down. Incubate for 15 minutes at 37˚C. | |||
| Yield | 300-500 μg. |
||
For making PCR fragment for in vitro transcription
Method 1: making T7 promoter + BGH tail
CMV-seq-F
GAGGTCTATATAAGCAGAGCTC
BGH-mRNA-R
GGTTCTTTCCGCCTCAGAAG
Method 2: making modified promoter AG + BGH tail
T7-mRNA-F1 (for pCDNA5 plasmid)
AAATTAATACGACTCACTATAAG
or T7-mRNA-F2 (for GOI in pCDNA5 and pCMV5-Frt plasmid vector)
ATTAATACGACTCACTATAAGGAGACCCAAG
BGH-mRNA-R
GGTTCTTTCCGCCTCAGAAG
with Flag Tag and polyA: use Flag-R
TTTTTTTTTTTTTTTCTCGAGTTACTTGTCGTCATCGTCTTTG
(CAAAGACGATGACGACAAGTAACTCGAGAAAAAAAAAAAAAAA)
PCR product sequence
ATTAATACGACTCACTATAAGGAGACCCAAG
<span style="color: rgb(37, 37, 37); font-family: sans-serif; font-size: 14px;">ATTAATACGACTCACTATAAGGAGACCCAAGCTGGCTAGC
GCTAGC CACCATG--GOI --ggatcc gactaCAAAGACGATGACGACAAGTAACTCGAGAAAAAAAAAAAAAAA</span>
Method 3: make T7 promoter + IRES+ BGH tail
first PCR
fragment 1
2711-IRES-F
ACGATGATAATATGGCCAGCTAGCCACCATGTTCGTCTTC
BGH-mRNA-R
GGTTCTTTCCGCCTCAGAAG
fragment 2
T7-mRNA-F3 (for plasmid with traditional T7 promoter and kozak seq, such as pMiniT2--IRES )
TAATACGACTCACTATAAGGAGG
IVT-IRES-R
CATGGTGGTGGCCATATTATC
2nd PCR
T7-mRNA-F3 (for plasmid with traditional T7 promoter and kozak seq, such as pMiniT2--IRES)
TAATACGACTCACTATAAGGAGG
TAATACGACTCACTATAAGGAGgccACCATG
GH-mRNA-R
GGTTCTTTCCGCCTCAGAAG
For GOI in pCMV5-Frt vector T7-mRNA-F3
TAATACGACTCACTATAAGGAGG
Method 4: make T7 promoter + polyA
For example: 2683 pJet1.2-SARS2-S1-1 PCR using
T7-mRNA-F3 TAATACGACTCACTATAAGGAGG
on fragment on pJet1.2
TAATACGACTCACTATAAGGAGG TAATACGACTCACTATAGGGAGgccAccATGAGAACTCAATTACCCCCTGCATACACTAATTCTTTCACACGTGGTGTTTATTACCCTGACAAAGTTTTCAGATCCTCAGTTTTACATTCAACTCAGGACTTGTTCTTACCTTTCTTTTCCAATGTTACTTGGTTCCATGCTATACATGTCTCTGGGACCAATGGTACTAAGAGGTTTGATAACCCTGTCCTACCATTTAATGATGGTGTTTATTTTGCTTCCACTGAGAAGTCTAACATAATAAGAGGCTGGATTTTTGGTACTACTTTAGATTCGAAGACCCAGTCCCTACTTATTGTTAATAACGCTACTAATGTTGTTATTAAAGTCTGTGAATTTCAATTTTGTAATGATCCATTTTTGGGTGTTTATTACCACAAAAACAACAAAAGTTGGATGGAAAGTGAGTTCAGAGTTTATTCTAGTGCGAATAATTGCACTTTTGAATATGTCTCTCAGCCTTTTCTTATGGACCTTGAAGGAAAACAGGGTAATTTCAAAAATCTTAGGGAATTTGTGTTTAAGAATATTGATGGTTATTTTAAAATATATTCTAAGCACACGCCTATTAATTTAGTGCGTGATCTCCCTCAGGGTTTTTCGGCTTTAGAACCATTGGTAGATTTGCCAATAGGTATTAACATCACTAGGTTTCAAACTTTACTTGCTTTACATAGAAGTTATTTGACTCCTGGTGATTCTT
CTTCAGGTTGGACAGCTGGTTAGTAACCGCTGAGCAATAACTAGCAAAAAAAAAA PCR Template Preparation
1. Cloning of IDT gene fragment into pMiniT2 or pJet1.2
2683 (DS1)
TAATACGACTCACTATAGGGAGgccAccATGAGAACTCAATTACCCCCTGCATACACTAATTCTTTCACACGTGGTGTTTATTACCCTGACAAAGTTTTCAGATCCTCAGTTTTACATTCAACTCAGGACTTGTTCTTACCTTTCTTTTCCAATGTTACTTGGTTCCATGCTATACATGTCTCTGGGACCAATGGTACTAAGAGGTTTGATAACCCTGTCCTACCATTTAATGATGGTGTTTATTTTGCTTCCACTGAGAAGTCTAACATAATAAGAGGCTGGATTTTTGGTACTACTTTAGATTCGAAGACCCAGTCCCTACTTATTGTTAATAACGCTACTAATGTTGTTATTAAAGTCTGTGAATTTCAATTTTGTAATGATCCATTTTTGGGTGTTTATTACCACAAAAACAACAAAAGTTGGATGGAAAGTGAGTTCAGAGTTTATTCTAGTGCGAATAATTGCACTTTTGAATATGTCTCTCAGCCTTTTCTTATGGACCTTGAAGGAAAACAGGGTAATTTCAAAAATCTTAGGGAATTTGTGTTTAAGAATATTGATGGTTATTTTAAAATATATTCTAAGCACACGCCTATTAATTTAGTGCGTGATCTCCCTCAGGGTTTTTCGGCTTTAGAACCATTGGTAGATTTGCCAATAGGTATTAACATCACTAGGTTTCAAACTTTACTTGCTTTACATAGAAGTTATTTGACTCCTGGTGATTCTTCTTCAGGTTGGACAGCTGGTTAGTAACCGCTGAGCAATAACTAGCAAAAAAAAAA
2. Make miniprep, send miniprep DNA out for sequencing (Genewiz, Macrogen)
3, PCR
1 ul of plasmid (50~100 ng)
1 ul of 10 uM T7-mRNA-F3 TAATACGACTCACTATAAGGAGG
1 ul 1 uM 2683-R2 2683-R2 TTACTTGTCGTCATCGTCTTTGTAGTCACCAGCTGTCCAACCTGAAG (CTTCAGGTTGGACAGCTGGTGACTACAAAGACGATGACGACAAGTAA)
1 ul 10 uM Flag-R
TTTTTTTTTTTTTTTCTCGAGTTACTTGTCGTCATCGTCTTTG
(CAAAGACGATGACGACAAGTAACTCGAGAAAAAAAAAAAAAAA)
21 ul H2O
+ 25 ul 2 X Phusion PCR mix (NEB)
PCR:
2 min at 95 oC
30 cycles of 95 oC 30 sec, 55 oC 30 sec, 72 oC 1 min
7 min at 72 oC.
Final Product 802 bp
TAATACGACTCACTATAAGGAGG
ccAccATGAGAACTCAATTACCCCCTGCATACACTAATTCTTTCACACGTGGTGTTTATTACCCTGACAAAGTTTTCAGATCCTCAGTTTTACATTCAACTCAGGACTTGTTCTTACCTTTCTTTTCCAATGTTACTTGGTTCCATGCTATACATGTCTCTGGGACCAATGGTACTAAGAGGTTTGATAACCCTGTCCTACCATTTAATGATGGTGTTTATTTTGCTTCCACTGAGAAGTCTAACATAATAAGAGGCTGGATTTTTGGTACTACTTTAGATTCGAAGACCCAGTCCCTACTTATTGTTAATAACGCTACTAATGTTGTTATTAAAGTCTGTGAATTTCAATTTTGTAATGATCCATTTTTGGGTGTTTATTACCACAAAAACAACAAAAGTTGGATGGAAAGTGAGTTCAGAGTTTATTCTAGTGCGAATAATTGCACTTTTGAATATGTCTCTCAGCCTTTTCTTATGGACCTTGAAGGAAAACAGGGTAATTTCAAAAATCTTAGGGAATTTGTGTTTAAGAATATTGATGGTTATTTTAAAATATATTCTAAGCACACGCCTATTAATTTAGTGCGTGATCTCCCTCAGGGTTTTTCGGCTTTAGAACCATTGGTAGATTTGCCAATAGGTATTAACATCACTAGGTTTCAAACTTTACTTGCTTTACATAGAAGTTATTTGACTCCTGGTGATTCTT
CTTCAGGTTGGACAGCTGGTGACTACAAAGACGATGACGACAAGTAACTCGAGAAAAAAAAAAAAAAA
MRTQLPPAYTNSFTRGVYYPDKVFRSSVLHSTQDLFLPFFSNVTWFHAIHVSGTNGTKRFDNPVLPFNDGVYFASTEKSNIIRGWIFGTTLDSKTQSLLIVNNATNVVIKVCEFQFCNDPFLGVYYHKNNKSWMESEFRVYSSANNCTFEYVSQPFLMDLEGKQGNFKNLREFVFKNIDGYFKIYSKHTPINLVRDLPQGFSALEPLVDLPIGINITRFQTLLALHRSYLTPGDSSSGWTAGDYKDDDDK
purify PCR fragment using Qiagen gel purification kit
Use 1 ug for IVT Purify PCR fragment using Qiagen gel purification kit
Use 1 ug for IVT

purify PCR fragment
use 1 ug for IVT
Add an EGFP tag to Flag tag Clone
[edit]EGFP PCR template
T7-EGFP-F
TAATACGACTCACTATAAGGAGACCATGGTGAGCAAGGGC
EGFP-R
tttttttTTTTTTTTTTTTTTTTACTTGTACAGCTCGTCC
(GGACGAGCTGTACAAGTAAAAAAAAAAAAAAAAaaaaaaa)
SARS2-EGFP fusion PCR template
clones should already have a Flag tag
to PCR out clone, such as 2686, 2689, 2690, 2691 original IDT fragments in pJet1.2
T7-mRNA-F4
TAATACGACTCACTATAAGGAGgccACCATG
Flag-EGFP-R
ctcctcgcccttgctcaccttgtcgtcatcgtctttgtag
and PCR EGFP from 2743 pLVX-Spike-IRES-EGFP
Flag-EGFP-F
ctacaaagacgatgacgacaaggtgagcaagggcgaggag
EGFP-R
tttttttTTTTTTTTTTTTTTTTACTTGTACAGCTCGTCC
(GGACGAGCTGTACAAGTAAAAAAAAAAAAAAAAaaaaaaa)
gel purified two fragments and combine
PCR again using
T7-mRNA-F4
EGFP-R
tttttttTTTTTTTTTTTTTTTTACTTGTACAGCTCGTCC
(GGACGAGCTGTACAAGTAAAAAAAAAAAAAAAAaaaaaaa)
purify and use it as template for IVT
transfection 293T cells with purified mRNA to see GFP expression
For amplifying DNA template from pCDNA5-FRT
[edit]T7-mRNA-F2
ATTAATACGACTCACTATAAGGAGACCCAAG
pCDNA5-mRNA-R
TTTTTTTTTTTTTTTTTTTTGGGGTCGAGGTGCCGTAAAG
(CTTTACGGCACCTCGACCCCAAAAAAAAAAAAAAAAAAAA)
Current Working Protocol
[edit]mRNA for transfection in vitro
[edit]Template preparation by PCR using 2X Taq PCR mix
| Clone # |
F Primer |
R Prime | PCR Length |
After PCR, take 5 ul to check purity on agarose gel. Purify the remaining PCR product using a spin column, elute DNA using 25 ul ddH2O, measure DNA concentration using nanodrop
mRNA as vaccine for animal study
[edit]Template preparation by PCR using 2X Taq PCR mix
| Clone # |
Name | Des | F Primer |
R Prime | PCR Length |
| 2684 |
DS2 | T7-IRES-DS2 |
T7-IRES-F or T7-F or IVT-T7-F |
Flag-R |
997 |
| 2838 | DS2 | T7-DS2 in pJet | T7-DS2-F |
Flag-R | 454 |
| 2831 |
DS2-fd | T7-DS2-fd in pJet | T7-DS2-F |
Foldon-R |
500 |
| 2823 | DS2 | T7-DS2 in pJet |
T7-DS2-F |
Flag-R |
454 |
| 2823 | DS2 SAM | T7-DS2 in pT7-VEE |
T7 or T7-VEE-F | Flag-R or VEE-pA-R |
8025 |
| 2831 | DS2-fd SAM | T7-DS2-fd in pT7-VEE | T7 or T7-VEE-F | Foldon-R or VEE-pA-R |
8065 |
| 2874 |
RBD-fd | Full length RBD fuse to fd, with polyA |
Flag-R TTTTTTTTTTTTTTTCTCGAGTTACTTGTCGTCATCGTCTTTG
(CAAAGACGATGACGACAAGTAACTCGAGAAAAAAAAAAAAAAA)
T7-DS2-F TAATACGACTCACTATAAGGAGGCCACCATGGTCAGACAAATC
Foldon-R tttttttttttttttttttttacaagaaggtcgacagcag
(ctgctgtcgaccttcttgtaaaaaaaaaaaaaaaaaaaaa)
VEE-pA-R GAATTAATTCCCCTCGAcgcgttctag
T7-VEE-F TAATACGACTCACTATAAGATGGGCGGCGCATGAGAG
After PCR, take 5 ul to check purity on agarose gel. Purify the remaining PCR product using a spin column, elute DNA using 25 ul ddH2O, measure DNA concentration using nanodrop


Use EMCV IRES to replace cap
[edit]PCR EMCV IRES from 959,961,960
T7 promoter: TAATACGACTCACTATAGGG
EMCV-IRES-R GGTGGTGGCCATATTATCATC
PCR from GOI 2684 pJet1.2-SARS2-S1-2
Spike RDB DS2
F: GATGATAATATGGCCACCACCCATATGGTCAGACAAATCGCTCCAG
R:
Make Self-amplifying mRNA (SAM) using VEE vector
[edit]PCR amplify
PCR making template from pCAGGS vector
[edit]for making mRNA PCR using
pCAGGS-T7
TAATACGACTCACTATAAGcattttggcaaggccaccatg
pCAGGS-pA
tTTTTTTTTTTTTTTTTTTTTTCATGGCAGCCAGCATATG
(CATATGCTGGCTGCCATGAAAAAAAAAAAAAAAAAAAAAa)
template: 2725 pCAGGS-SARS2-RBD or 2754 pCAGGS-SARS2-S-6H
mRNA from 2874 pCDNA5-frt-SARS2-RBD-fd
[edit]RBD trimer, signal peptide, Codon optimized, 190 poly A tail
1, PCR
[edit]primers
T7-mRNA-F2 ATTAATACGACTCACTATAAGG AGACCCAAG
2874-R
AACGGGCCCTCTAGActcgAGAATTAATTCCCCTCGAcgc
(gcgTCGAGGGGAATTAATTCTcgagTCTAGAGGGCCCGTT)
extension time 1 min
precipitate using isopropanol
optionally, remove extra nt from 3' using PacI or Xho1
2, Digestion plasmid DNA with PacI, or Xho1 or MluI or XbaI
[edit]PacI digested template has many large RNA or DNA bands,
For T7-VEE, PacI and XhoI produces 3´overhang, so they are not good for linearization, if PacI or Xho1 is to be used, DNA template must be blunted by adding DNA Polymerase I Large (Klenow) Fragment at a concentration of 5u/μg and incubate the reaction mixture for 15 minutes at 22°C.
For digestion of 36 ug of T7-VEE plasmid in 200 μL
36 μL T7-VEE Plasmid (1 μg/μL)
20 μL CutSmart Buffer (10X)
139 μL Sterile distilled water
5 μL MluI-HF (10,000 units/mL, NEB)
Incubate digestion mixtures at 37°C for 3 hours.
Add the same volume (200 μL) of phenol/chloroform/isoamyl alcohol (25:24:1) to each eppendorf tube containing the digestion mixture.
Vortex the eppendorf tubes for 60 seconds. Centrifuge at high speed (14,000 rpm) for 5 minutes.
Transfer each aqueous phase (top layer) into new eppendorf tubes.
Note: T7-VEE plasmid is a large sized DNA and may be trapped in the interphase layer with the proteins. When the interphase is sticky (DNA is trapped), collect the interphase layer along with the aqueous layer.
To remove the excess phenol, add an equal volume (200 μL) of chloroform/isoamyl alcohol (24:1) to each eppendorf tubes.
Vortex the eppendorf tubes for 60 seconds and centrifuge at high speed (14,000 rpm) for 5 minutes.
Transfer each aqueous phase (top layer) into new eppendorf tubes.
Add 1/10 volume (20 μL) of 3 M sodium acetate, pH 5.2 (this can be found in the RiboMAX Large Scale RNA Production kit) and 2.5 X volume (500 μL) of 100% ethanol to each eppendorf tubes.
Vortex and place at -80°C for 5 minutes or store at -20°C until use.
Note: If there are no plans to proceed to RNA synthesis on the same day, stop at this step. DNA will be stable for several months at -20°C.
Centrifuge at high speed (14,000 rpm) for 10-15 minutes at 4°C to pellet the DNA plasmid. Remove the ethanol carefully with a pipet tip, being careful not to dislodge the DNA pellet.
Gently add 1 mL 70% ethanol to wash the DNA pellet. Centrifuge at high speed at 4°C for 1 minute. Remove the ethanol carefully with a pipet tip, being careful not to dislodge the RNA pellet.
Centrifuge shortly, and carefully remove any visible residual ethanol with a P20 pipette. Drying the DNA pellet is not necessary. In general, approximately 20% of plasmid may be lost during purification.
Resuspend the DNA pellet in a small volume (15~20 μL, depending on starting amount) of DNase & RNase-free water before measuring the concentration.
Measure the DNA concentration of each linearized DNA plasmid. Adjust the concentration to 1 μg/μL.
mRNA analysis by agarose
| Lane | Desc | Note |
| 1 | PacI digested plasmid, large scale | need to remove DNA using DNase I |
| 2 | PacI digested plasmid, small scale |
|
| 3 | Xho1 digested plasmid, small scale | |
| 4 | PCR template, small scale | |
| 5 | DNA ladder, NEB, 1 Kb | |
| 6 | PCR template 1, precipitate with isopropanol + glycogen |
|
| 7 | PCR template 2, precipitate with isopropanol |

mRNA made from 6 + 7 combined, 4 tubes of 40 ul reaction from template 6, 4 tubes of 40 ul reaction from template 7, purified using Qiagen RNeasy miniprep kit

CleanCap is used for easy one -step reaction, while enzymatic capping method requires three-step reaction but it minimizes the IFN responses by combination with phosphatase treatment
High yield Production of mRNA
[edit]10 mL transcription reaction mixture
30 mM Tris (pH 8.1 at 37°C), 15 mM Mg2+ , 10 mM dithiothreitol (DTT), 2 mM spermidine, 0.01% (v/v) Triton X-100, 4 mM each NTP, 1 mL of PCR-generated DNA template, and 0.1 mg/mL of T7 RNAP
optimal concentration of Mg2+ is 15 ~45 mM
After 2.5 h of incubation at 37°C, pyrophosphate, which forms during in vitro transcription reaction, was pelleted down by centrifugation, and additional Mg2+ should be added to the reaction.
The reaction continued till 5 h.
Size-exclusion FPLC based RNA purification
RNA sample preparation for chromatographic purification
Transcription mixtures for the S-box RNA were heated to 90°C for 5 min, immediately transferred to ice for 5 min and then pelleted down (centrifuged at 16, 000g for 10 min). The supernatant was collected and mixed as 1: 4 with Tris-EDTA buffer containing 8 M urea and 100 mM NaCl before loading onto size-exclusion FPLC (SEC) columns. Sample pre-load treatment was operated to remove the T7 RNAP, residual pyrophosphates and aggregates formed during in vitro transcription.
Denaturing chromatographic purification
The glass columns (890/10 mm) and (xk16/1000 mm) were packed according to manufacturer’s specifications with gel filtration matrix (Superdex-75, Superdex-200) and equilibrated in a cold room (4°C) with several column volumes (CV) of specified buffer. The transcription mixture from the previous step was loaded directly onto the equilibrated SEC column, and the chromatography was performed at 1 mL/min flow rate, collecting 2 mL fractions. Fractions were analyzed by denaturing urea-PAGE. S-box RNA containing fractions were combined and concentrated to 0.5 mL using Millipore centrifugal filter units (EMD Millipore), washed twice with the appropriate buffer for chemical probing analysis. All experiments were performed at 4°C. The efficiency of proposed purification system was confirmed through SHAPE experiments for S-box RNA.
https://www.targetmol.com/compound/mrtx-1719
Transfection
[edit]Thaw mRNAs were thawed on ice and denatured at 65°C for 3min
Incubation at room temperature for 45min.
1. Lipofectamine 3000
- On day 0, HEK 293T cells are seeded at a concentration of 5 × 10⁵ cells/well in a 6-well plate to ascertain 70–90 % of confluency on the day of transfection (day 1).
- For transfection, a ratio of 2 μl of Lipofectamine® reagent to 1 μg of RNA is used. Cells are transfected with 1 and 2 μg of mRNA. Transfection is performed in duplicates.
- On the day of transfection, an appropriate volume of Lipofectamine® reagent is prepared in Opti-MEM® medium and incubated at room temperature for 5 min.
- 1 and 2 μg of mRNA are diluted in Opti-MEM® medium. If required, pre-dilute mRNA to a concentration of 0.5 μg/μl.
- Mix Lipofectamine® reagent and Opti-MEM®-diluted mRNA gently in a total volume of 500 μl at a 1:1 ratio (v/v) and incubated at room temperature for 20 min.
- Wash cells twice with 500 μl DMEM medium containing glutamine but without FBS. Add 2 ml of DMEM to the cells.
- Add 500 μl of mRNA–lipid complexes to each well. Mix gently.
- Incubate cells at 37 °C in a CO2-incubator for 4–6 h.
- After the incubation, replace the medium with 2 ml of fresh DMEM medium containing 10 % FBS.
2. TT3 nanoparticles
lipid:DOPE:cholesterol:DMG-PEG2000 at a molar ratio of 20:30:40:0.75 (https://www.ncbi.nlm.nih.gov/pmc/articles/PMC4869688/)
3. Protamine as an immunostimulatory component
Protamine. Pharmaceutical Protamine is available at two concentrations: 1000 (1 ml neutralizes 1000 IU of heparin) or 5000 (1 ml neutralizes 5000 IU of heparin). To generate particles of 130 nm, we use Protamine Ipex 5000 from MEDA Pharmaceuticals (Protamine hydrochloride 5000 IU/ml). The Protamine is stored at 4 °C
Dissolve the protamine in Ringer lactate solution.
While stirring, slowly add the protamine-Ringer lactate solution half of the mRNA until you reach a weight ratio of mRNA and protamine of 2:1.
Stir the solution for another 10 min to ensure the formation of stable complexes.
Add the remaining half of free mRNA and stir briefly.
Adjust the final concentration of the vaccine with Ringer lactate solution.
Carry out all procedures at room temperature unless otherwise specified.
This step is performed under laminar flow using sterile equipment.
- Prepare a 0.5 mg/ml Protamine solution (see Note 5) by diluting 10 μl of Protamine 5000 in 280 μl of water.
- Prepare a 0.5 mg/ml RNA solution (see Note 5) by diluting the RNA stock with the appropriate amount of water.
- Add an equal amount of Protamine solution to the RNA solution (see Note 6), and pipet quickly up and down at least ten times (see Note 7).
- Leave the solution at room temperature for 10 min (see Note 8). 1. Dilute 40 μl of the Protamine–RNA formulation (containing 10 μg of RNA and 10 μg of Protamine) with 1 ml of water (see Note 9).
Set the viscosity of the analyzer to 0.89 (select within the software setting either measurement in water or a viscosity of 0.89). As shown in Fig. 1, particles made in the conditions described above are approximately 130 nm in size with a polydispersity index (PDI) of less than 0.3 (see Note 10).
Protamine sulfate, such as Valeant Protamine, gave identical results.
Freezing Protamine solutions may jeopardize its functionality as far as production of immunostimulating RNA polyplex is concerned. Pharmaceutical Protamine should be stored at 4 °C as indicated on the package.
Any RNA (oligonucleotide, long uncapped and/or non-polyadenylatedmRNA, or capped mRNA) can be used to generate the 130 nm particles
Nonobody mRNA
[edit]HiScribe T7 ARCA mRNA kit (with tailing; NEB; #E2060S). PCR product should be purified and run on an agarose gel to confirm amplicon size prior to its use as a template in the T7 ARCA mRNA transcription reaction. Generally, 0.1–0.5 mg of PCR fragments can be used in a 20-mL in vitro transcription reaction. After the preparation of PCR templates, standard mRNA synthesis with modified nucleotides should be started with thawing the necessary kit components, mixing, and pulse spinning in a microfuge to collect solutions to the bottoms of the tubes. The reaction was assembled at room temperature in the following order: nuclease-free water (to 20 uL), 2 x ARCA/NTP mix (10 uL), 5mCTP (10 mM, 2.5 uL), pseudo-UTP (10 mM, 2.5 uL), template DNA (1 mg), and T7 RNA polymerase mix (2 uL); mixed thoroughly; and pulse spun in a microfuge. Incubation was at 37oC for 30 min. DNase treatment to remove template DNA included adding 2 uL of DNase I, mixing well, and incubating at 37 oC for 15 min. poly(A) tailing starts with setting up the tailing reaction as below. Standard tailing reaction volume was 50 uL: H2O (20 uL), IVT reaction (20 uL), 10 X poly(A) polymerase reaction buffer (5 uL), and poly(A) polymerase. It was mixed thoroughly, pulse spun in a microfuge, and incubated at 37oC for 30 min. Synthesized mRNA can be purified by LiCl precipitation, phenol/chloroform extraction, followed by ethanol precipitation. To the 50-mL tailing reaction, 25 mL LiCl solution was added and mixed well. Incubation at -20 oC for 30 min was completed, as was centrifugation at 4oC for 15 min at top speed to pellet the RNA. The supernatant was removed carefully, the pellet was rinsed by adding 500 mL of cold 70% ethanol and centrifuged at 4oC for 10 min, ethanol was removed carefully, the tube was spun briefly to bring down any liquid on the wall, residual liquid was removed carefully using a sharp tip, the pellet was air dried, the mRNA was resuspend in 50 mL of 0.1 mM EDTA or a suitable RNA storage solution, and the RNA was heated at 65oC for 5– 10 min to completely dissolve the RNA and mixed well.
LNP
[edit]LNP were prepared by mixing ethanol and aqueous phase at a 1:3 volumetric ratio in a microfluidic device, using syringe pumps as previously described (Chen, Love et al., 2012). The ethanol phase was prepared by solubilizing a mixture of ionizable lipidoid cKK-E12, 1,2-dioleoyl-sn-glycero-3-phosphoethanolamine (DOPE, Avanti), cholesterol (Avanti), and 1,2-dimyristoyl-sn-glycero-3-phosphoethanolamine-N-[methoxy-(polyethyleneglycol)-2000] (ammonium salt) (C14-PEG 2000, Avanti) at a molar ratio of 35:16:46.5:2.5 The aqueous phase was prepared in 10 mM citrate buffer (pH 3) with either trastuzumab m 164 RNA or luciferase mRNA (Firefly luciferase mRNA, TranslateBio). LNPs were dialyzed against PBS in a Slide-A-Lyzer™ G2 Dialysis Cassettes, 20,000 MWCO (Thermo Fisher) for 2h at RT. The concentration of mRNA encapsulated into LNPs nanoparticles was analyzed using Quant-iT RiboGreen assay (Thermo Fisher), according to manufacturer’s protocol. The efficacy of mRNA encapsulation into LNPs was calculated by comparing measurements in the absence and presence of 1% (v/v) Triton X- 100. Nanoparticle size, polydispersity (PDI), and ζ-potential were analyzed by dynamic light scattering (DLS) using Zetasizer Nano ZS (Malvern Instruments, Worcestershire, UK). LNP hydrodynamic diameters are reported in the volume weighting mode and are an average of three independent measurements (Supplemental Table 5).
RNA capping using Vaccinia Capping Enzyme and Cap 2´-O-Methyltransferase
[edit]IVT reactions: 1 µM primers, 40 mM Tris pH 8, 5 mM DTT, 1 mM spermidine, 0.01% Triton X-100, 4 mM each of NTP in the RNA sequence, 0.2 µM T7 RNA polymerase, 20–60 mM MgCl2.
RNA purification using anion-exchange chromatography
[edit]RNA from IVT can be purified using anion exchange chromatography on a 22 × 250 mm DNAPac PA100 column (Dionex) operated at 80°C. https://www.thermofisher.com/order/catalog/product/043011#/043011
The column is run in buffers containing 20 mM Tris pH 8.0 and 5 M Urea (pH adjusted to 8.0 at room temperature). RNA of different lengths is eluted with a NaCl gradient. Salt gradients can be designed according to the respective RNA lengths to be separated. Short RNAs (2–10 nt) typically elute between 50 and 200 mM NaCl while longer constructs (60–80 nt) elute at 300–400 mM NaCl. RNA containing fractions can be analyzed using urea-PAGE, pooled and precipitated as below. After precipitation, RNAs can be desalted using a prepacked PD-10 Desalting column (GE Healthcare) and subsequently concentrated using a speedvac device.
RNA precipitation
[edit]RNA can be precipitated with 0.3 M NaOAc pH 5.2 and 0.7 volumes of isopropanol at −20°C for at least 2 h. The reaction is then centrifuged at 9000g and −5°C for 1 h and the pellet is washed with cold 75% ethanol and centrifuged again for 30 min. The resulting pellet is air dried and resuspended in the desired buffer for further applications. For short RNAs (2–10 nt), precipitation can be carried out with 0.2 M NaCl and 3.5 volumes of EtOH (replacing the NaOAc and isopropanol) and a precipitation time of 12 h
RNA capping
[edit]Capping reactions is performed in buffer containing 50 mM Tris, pH 8, 5 mM KCl, 1 mM MgCl2, and 1 mM DTT with 20 µM RNA. 0.5 mM GTP, 0.1 mM SAM, and capping enzyme according to the previously determined substrate to enzyme ratio is added. The reaction is incubated at 37°C and prepared for analysis by Urea-PAGE by adding an equal volume of sample buffer containing 8 M Urea, 20 mM EDTA, 2 mM Tris pH 8, and Bromophenol blue and Xylene Cyanol.
For cap-1: cap-1 RNA, Vaccinia MTase (NEB) is added to the capping reaction
RNA promoter sequence Promoter sequence T7 promoter Φ6.5 (for start with G) CGTAATACGACTCACTATAGG T7 promoter Φ2.5 (for start with A) CGTAATACGACTCACTATTAG
Cas13a
[edit]In vitro transcription of Cas13a. Plasmids were linearized with Not-I HF (New England Biolabs) overnight at 37 °C. Linearized templates were purified by sodium acetate (Thermo Fisher Scientific) precipitation and rehydrated with nuclease-free water. In vitro transcription was performed overnight at 37 °C using the HiScribe T7 Kit (NEB) following the manufacturer’s instructions (N1-methyl-pseudouridine modified). The resulting RNA was treated with DNase I (Aldevron) for 30 min to remove the template, and it was purified using lithium chloride precipitation (Thermo Fisher Scientific). The RNA was heat denatured at 65 °C for 10 min before capping with a Cap-1 structure using guanylyl transferase and 2′-O-methyltransferase (Aldevron). mRNA was then purified by lithium chloride precipitation, treated with alkaline phosphatase (NEB) and purified again. mRNA concentration was measured using a Nanodrop. mRNA stock concentrations were 1–3 mg ml−1. Purified mRNA products were analyzed by gel electrophoresis to ensure purity. crRNA guides were purchased from Integrated DNA Technologies or GenScript https://www.aldevron.com/hubfs/content/Documents/ald-IVT-enzyme-tech-guide-090010916.pdf
Enzymes for mRNA research
[edit]https://www.ncbi.nlm.nih.gov/pmc/articles/PMC4986899/
T7 RNA Polymerase 9135-130MCG 1
2598 pET29-Turbo-T7-RNAPo
E coli Inorganic Pyrophosphatase 9132-350MCG 1
2259 need to do cloning
DNase I 9136-70MCG 1
202 pET28-DNase1 need to subclone into pMBP vector
Ribonuclease Inhibitor 9134-180MCG 1
pT7A30-RNaseInhibitor Sus scrofa (wild pig) RNase Inhibitor 1499
Guanylyltransferase 9131-300MCG 2
2'-O-Methyltransferase 9130-1450MCG 2 An E. coli strain that carries the gene for the Vaccinia mRNA Cap 2´-O-Methyltransferase. Poly(A) Polymerase 9133-430MCG 2
Ribonuclease Inhibitor 9134-660MCG 2
Guanylyltransferase Catalyzes addition of 7-methylguanylate cap structure (Cap 0) to the 5’ end of RNA in three steps:
2’-O-Methyltransferase Reactions 1 & 2: Converts Cap 0 to Cap 1 by adding a methyl group to the ribose 2’-OH of the first nucleotide adjacent to the cap structure. Activity can be measured using the EPIgeneousTM Methyltransferase Assay (Cisbio).
Reaction 3: Conversion of an RNA substrate pppN(pN)n to GpppN(pN)n with increasing amounts of enzyme can be visualized by gel electrophoresis (uncapped, lower band; capped, upper band).
Methyl transfer from SAM to GpppN(pN)n producing SAH and Cap 0 mRNA can be measured using the EPIgeneousTM Methyltransferase Assay (Cisbio).
https://www.aldevron.com/products/ivt-enzyme-kits
IVT Buffer: https://www.ncbi.nlm.nih.gov/pmc/articles/PMC306405/pdf/nar00265-0210.pdf
40 mM Tris-HCl (pH 8.1 at 37°C), 1 mM spermidine, 5 mM dithiothreitol, 50 ug/ml bovine serum albumin, 0.01% (vlv) Triton X-100 and 80 mg/ml polyethylene glycol (8000 MW)
https://www.neb.com/gmp/gmp-grade-reagents-for-rna-synthesis
Inorganic pyrophosphatase https://www.neb.com/products/m0361-pyrophosphatase-inorganic-e-coli PPase is prepared from an E. coli strain containing a clone of the E. coli inorganic pyrophosphatase gene.
DNaseI : https://www.neb.com/products/m0303-dnase-i-rnase-free An E. Coli strain that carries an MBP fusion clone of Bovine Pancreatic DNase I.
Flp mRNA
[edit]PCR using these two primers
Template; 2895 T7-VEE-NLS-Flp
T7-HBG-F
TAATACGACTCACTATAAGGACATTTGCTTCTGACACAAC
VEE-pA-R GAATTAATTCCCCTCGAcgcgttctag
sequence
T7 (modified)
TAATACGACTCACTATAAGG
ACATTTGCTTCTGACACAAC
TGTGTTCACTAGCAACCTCAAACAATT
Kozak
GCCACC
ATGGCCCCCAAGAAAAAGCGGAAAGTGATGAGCCAGTTCGACATCCTGTGCAAGACCCCCCCCAAGGTGCTCGTGCGGCAGTTCGTGGAAAGATTCGAGAGGCCCAGCGGCGAGAAGATCGCCTCTTGTGCCGCCGAGCTGACCTACCTGTGCTGGATGATCACCCACAACGGCACCGCCATCAAGCGGGCCACCTTCATGAGCTACAACACCATCATCAGCAACAGCCTGAGCTTCGACATCGTGAACAAGAGCCTGCAGTTCAAGTACAAGACCCAGAAGGCCACCATCCTGGAAGCCAGCCTGAAGAAGCTGATCCCCGCCTGGGAGTTTACCATCATCCCCTACAACGGCCAGAAGCACCAGAGCGACATCACCGATATCGTGTCCAGCCTGCAGCTGCAGTTCGAGAGCAGCGAGGAAGCCGACAAGGGCAACAGCCACAGCAAGAAGATGCTGAAGGCCCTGCTGAGCGAGGGCGAGAGCATCTGGGAGATCACAGAGAAGATCCTGAACAGCTTCGAGTACACCAGCCGGTTCACCAAGACCAAGACCCTGTACCAGTTCCTGTTCCTGGCCACCTTTATCAACTGCGGCCGGTTCTCCGACATCAAGAACGTGGACCCCAAGAGCTTCAAGCTGGTGCAGAACAAGTACCTGGGCGTGATCATTCAGTGCCTCGTGACCGAGACAAAGACCAGCGTGTCCCGGCACATCTACTTTTTCAGCGCCAGAGGCCGGATCGACCCCCTGGTGTACCTGGACGAGTTCCTGAGAAACAGCGAGCCCGTGCTGAAGAGAGTGAACCGGACCGGCAACAGCAGCTCCAACAAGCAGGAATACCAGCTGCTGAAGGACAACCTCGTGCGGTCCTACAACAAGGCCCTGAAGAAAAACGCCCCCTACCCCATCTTCGCCATTAAGAACGGCCCCAAGTCCCACATCGGCCGGCACCTGATGACCAGCTTTCTGAGCATGAAGGGCCTGACAGAGCTGACCAACGTCGTGGGCAATTGGAGCGACAAGAGGGCCTCTGCCGTGGCCAGAACCACCTACACCCACCAGATCACAGCCATCCCCGACCACTACTTCGCCCTGGTGTCTCGGTACTACGCCTACGACCCCATCAGCAAAGAGATGATCGCCCTGAAGGACGAGACAAACCCCATCGAGGAATGGCAGCACATCGAGCAGCTGAAGGGCAGCGCCGAGGGCAGCATCAGATACCCTGCCTGGAACGGCATCATCTCCCAGGAAGTGCTGGACTACCTGAGCAGCTACATCAACCGGCGGATCTAG
Not1
GCGGCCGC
AAGCTGCTTACATAGAACTCGCGGCGATTG
Sph1
GCATGC
CGCCTTAAAATTTTTATTTTATTTTTtCTTTTCTTTTCCGAATCGGATTTTGTTTTTAATATTTCAAAAAAAAAAAAAAAAAAAAAAAAACGCGAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAttaatAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAttaatAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAA
PacI
ttaattaa
Xba1
tctaga
Mlu1
acgcgT CGAGGGGAATTAATTC
(VEE-pA-R GAATTAATTCCCCTCGAcgcgttctag )
https://www.targetmol.com/compound/mrtx-1719
EGFP mRNA Protocol
[edit]to make template for Cleancap by PCR
T7AG-HBG-F GTTAATACGACTCACTATAagGTCTTCTGGTCCCCACAGAC--> this is for cleancap
VEE-pA-R GAATTAATTCCCCTCGAcgcgttctag
T7-HBA2-F is not as good as T7AG-HBG-F
Template: 2885 T7-VEE-EGFP
(F primer T7AG-HBG-F GTTAATACGACTCACTATAagGTCTTCTGGTCCCCACAGAC- for cleancap)
sequence, 1113 bp
GTTAATACGACTCACTATAag TCTTCTGGTCCCCACAGACTCAGAGAGAACCC Kozak gccacc
EGFP atggtgTCTaagggcgaggagctgttcaccggggtggtgcccatcctggtcgagctggacggcgacgtaaacggccacaagttcagcgtgtccggcgagggcgagggcgatgccacctacggcaagctgaccctgaagttcatctgcaccaccggcaagctgcccgtgccctggcccaccctcgtgaccaccctgacctacggcgtgcagtgcttcagccgctaccccgaccacatgaagcagcacgacttcttcaagtccgccatgcccgaaggctacgtccaggagcgcaccatcttcttcaaggacgacggcaactacaagacccgcgccgaggtgaagttcgagggcgacaccctggtgaaccgcatcgagctgaagggcatcgacttcaaggaggacggcaacatcctggggcacaagctggagtacaactacaacagccacaacgtctatatcatggccgacaagcagaagaacggcatcaaggtgaacttcaagatccgccacaacatcgaggacggcagcgtgcagctcgccgaccactaccagcagaacacccccatcggcgacggccccgtgctgctgcccgacaaccactacctgagcacccagtccgccctgagcaaagaccccaacgagaagcgcgatcacatggtcctgctggagttcgtgaccgccgccgggatcactctcggcatggacgagctgtacaagtaa Not1 GCGGCCGC AAGCTGCTTACATAGAACTCGCGGCGATTG
Sph1
GCATGC
CGCCTTAAAATTTTTATTTTATTTTTtCTTTTCTTTTCCGAATCGGATTTTGTTTTTAATATTTCAAAAAAAAAAAAAAAAAAAAAAAAACGCGAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAttaatAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAttaatAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAAA
PacI
ttaattaa
Xba1
tctaga
Mlu1
acgcgT CGAGGGGAATTAATTC
(R primer VEE-pA-R GAATTAATTCCCCTCGAcgcgttctag )
Cy5-UTP
[edit]
IVT: Use 1 ul cy5-UTP for 80 ul reaction, 4 ul of buffer , 4 ul of each NTP, 4 ul enzyme, react 24 hours
25ul https://www.apexbt.com/cy5-utp.html, 25ul (10mM) $165.00, Cy5-UTP Catalog No. B8333
Cyanine https://www.perkinelmer.com/product/cyanine-5-utp-nel583001ea Cyanine 5-UTP, 100 nmol, NEL583001EA 577.00 USD
ENZ-42506 https://www.enzolifesciences.com/ENZ-42506/cyanine-5-utp-enhanced/ ENZ-42506 250 nmol 344.00 USD
PK-PFCY5-UTP-10 https://promocell.com/product/cy5-utp/ PK-PFCY5-UTP-10 10 µl $ 299.00
https://www.trilinkbiotech.com/cyanine-5-aminoallyluridine-5-triphosphate.html
PA55026Cy5-UTP 100 nmol 598.00 USD
Cyanine https://www.scbt.com/p/cyanine-5-utp Cyanine 5-UTP, 250 nmolsc-499469, 250 nmol $316.00


NEB Protocol:
[edit]4 ul of each: 100 mM ATP, GTP, UTP, CTP, 3.2 ul of 100mM Cleancap, 4 ul of 10X Buffer, 40 ul of 2885 PCR product (2 ug total), 1 ul of 10mM Cy5-UTP (santa cruz), 5 ul of T7 RNA Polymerase Mix, Water to 80 ul, reaction for 24 hours.
Purify using RNAeazy spin column
Yield: 150 ul of 1.7 ug/ul, 50 ul of 816ng/ul
Total yield: 255 ug + 40 ug= 295 ug
Lucigen AmpliScribe High Yield Transcription Kits Protocol
[edit]x µL Nuclease-Free Water,
Sterile 1 µg linearised template DNA with T7 promoter
2 µL AmpliScribe T7 10X Reaction Buffer
1.5 µL 100 mM ATP
1.5 µL 100 mM CTP
1.5 µL 100 mM GTP
1 µL 100 mM UTP
6 µL 4 mM Cy5-UTP
2 µL 100 mM DTT
0.5 µL RiboGuard RNase Inhibitor
2 µL AmpliScribe T7 RNA Polymerase
20 µL Total reaction volume
Incubate at 37 °C for 2 hours